
Tuesday, August 7, 2012
Cohera Medical Completes Enrollment of U.S. Clinical Trial of TissuGlu® Surgical Adhesive
PITTSBURGH, Aug. 7, 2012 /PRNewswire/ -- Cohera Medical, Inc.®, a leading innovator and developer of absorbable surgical adhesives and sealants, today announced the completion of enrollment of its prospective, multicenter, randomized clinical trial for its lead product, TissuGlu® Surgical Adhesive inthe United States.
The study is evaluating the effectiveness of TissuGlu on the reduction of wound drainage and post-surgical complications in abdominoplasty surgeries. The study included 150 patients enrolled at five sites throughout the U.S., including Atlanta, Baltimore, Charlotte, St. Louis, and Washington DC.
"We are delighted to complete enrollment in this critical trial for TissuGlu, which has the potential to eliminate wound drainage issues from large flap procedures," said Dr. Joseph Hunstad of the Hunstad-Kortesis Center for Plastic Surgery, Charlotte, NC. "The trial results so far are very promising for both the surgeons and the patients."
Completion of enrollment of the study marks a significant milestone in Cohera Medical's progression of its lead product toward commercialization in the United States. The Company will use the results from the study to submit a PMA application for TissuGlu with the FDA. There are currently no medical devices approved by the FDA, or in pivotal clinical trials, for a synthetic adhesive indicated for approximation of tissues in large flap surgeries.
"We are extremely pleased to have reached this very significant milestone in our clinical study," saidPatrick Daly, President and Chief Executive Officer of Cohera Medical. "It is a testament to the surgeons and the Cohera team to enroll 150 patients in 90 days. We are encouraged by the positive feedback from the TissuGlu investigators and look forward to our work with the FDA to make TissuGlu available to surgeons and patients throughout the U.S."
Cohera Medical received CE Marking approval for TissuGlu and began selling product to hospitals and surgeons in Germany in September 2011. To date, TissuGlu has been used successfully in over 500 surgical procedures in Germany by leading plastic and reconstructive surgeons.
Currently, most patients who undergo abdominoplasty procedures and other large flap procedures require the insertion of drains to remove fluids that accumulate under the skin at the surgical site. In some cases, drainage is inadequate and the excess fluid accumulation (seroma) requires additional procedures for removal. TissuGlu adheres the tissue flap created during the procedure to the underlying tissue, helping to reduce the fluid that can accumulate in the space and ultimately reducing the time to drain removal. The use of TissuGlu may shorten the length of time that drains need to be in place, leading to a more comfortable recovery and a quicker return to normal activity for patients.


Thursday, July 19, 2012
Sysmex and Siemens Renew Global Partnership for Hemostasis Solutions and Services
President and CEO: Hisashi Ietsugu) and Siemens Healthcare Diagnostics (HQ: NY, USA; CEO: Michael Reitermann) have announced the signing of a five-year global supply, distributorship, sales and service contract extension under which clinical laboratory customers around the world can retain access to the largest portfolio of innovative hemostasis instruments and reagents through 2018. The companies, which began partnering in 1995, also agreed to continue joint product development activities, ensuring customer access to new hemostasis technology. The signing occurred during the 2012 AACC and ASCLS Annual Meetings and Clinical Lab Expo in Los Angeles, California.
The large assortment of hemostasis products covered in this latest agreement is used by clinical laboratories to test for blood disorders in patients and to monitor patients on blood-thinning medication. To date, Sysmex and Siemens have released to market a variety of advanced solutions, including the recently launched Sysmex CS-5100*(1), the top-end model in Sysmex's CS hemostasis analyzer series, the Sysmex® CA-600 Systems*(1), a new line of compact, fully automated hemostasis analyzers, as well as several new Siemens INNOVANCE™ reagents.

"We are pleased to extend our longstanding relationship with Siemens Healthcare Diagnostics," said Hisashi Ietsugu, President and CEO of Sysmex Corporation. "Our partnership in the field of hemostasis combines the strength of two leading diagnostics companies whose joint expertise offers best-in-class solutions to the laboratory. We remain committed to further expand our offerings with enhanced clinical utility and workflow efficiency."
"Our shared goal to deliver high quality products and services that address our customers' needs is a key reason for the success of our 17-year relationship," said Michael Reitermann, CEO, Siemens Healthcare Diagnostics. "Sysmex and Siemens remains a reliable partnership for customers worldwide amidst an evolving hemostasis market. Together, we will continue offering customers the broad portfolio of hemostasis solutions they have come to expect from us and we will also focus on delivering next-generation technologies going forward."
Sunday, July 15, 2012
It's Perclot AMP.......No, It's Endoclot AMP

1.5 Product Applicators, Etc. SMI agrees to promptly notify CryoLife of all improvements to applicators, tips and other accessories included within or used in connection with the Products, including all new applicators, tips and other accessories. All such improvements and any such new applicators, tips or accessories to the Products shall be included within the Products and the Parties shall adjust the catalog of Products to reflect these new products, with any transfer price to be negotiated in good faith, but based solely on costs to SMI for such improvements and/or new applicators, tips and accessories. The Parties agree that the Endoscopic applicator system used for powder delivery via gastrointestinal endoscope, as further described on Schedule 2.1 is not an improvement or new applicator and is not included in this Agreement.
2. Distribution
2.1 Limitations on SMI Activities. During the term of this Agreement SMI agrees (i) to sell the Products exclusively to CryoLife for use in Permitted Clinical Applications within the Territory, (ii) to refrain from selling or licensing any Products to any Existing Distributor or Third Party for sale or distribution in Permitted Clinical Applications within the Territory, (iii) to refrain from directly or indirectly marketing, promoting, or encouraging any Third Party to market, promote or Distribute the Products for any of the Permitted Clinical Applications within the Territory, (iv) to refrain from licensing or transferring any AMP™ Technology to any Third Party within the Territory for the purpose of manufacturing any Products upon terms or conditions that would enable or allow such Third Party to sell any Products for Permitted Clinical Applications within the Territory. In addition, SMI agrees that it shall refrain until January 1, 2015 from (A) directly, or indirectly selling, permitting to sell, market, promote or encouraging third parties to sell, permit to sell, market or promote any Competitive Product (defined below) for any Permitted Clinical Application within the Territory or (B) licensing or transferring to any Third Party technology that would enable or allow any Third Party to manufacture any Competitive Product within the Territory. The provisions of the foregoing sentence shall be deemed further modified so that SMI may only take the actions described therein if SMI complies with Section 1.4 (and therefore CryoLife does not match the right of first refusal set forth therein). As used herein, “Competitive Product” means any powdered absorbable surgical hemostat that is intended for or could be used for a Permitted Clinical Application. The foregoing limitations do not apply to sales by SMI of those products described on Schedule 2.1.
Read the full document HERE

.
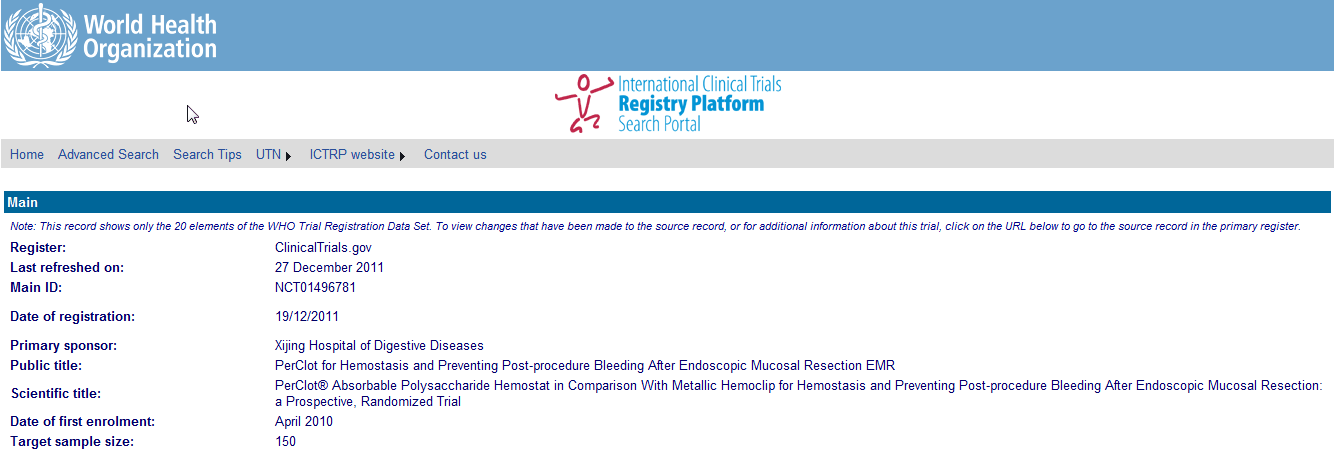
Now Cryolife and their partners face the interesting prospect of Chinese made Perclot AMP particles meeting them in the EU re-branded as "Endoclot".
As can be seen HERE the original Chinese Xijing Hospital trial changes for the use of AMP show the changes from perclot to "endoclot", but the endoclot webpage discusses AMP particles....see HERE.
The CRY Q2 Earnings Conference Call is scheduled for 7/31/2012.
Click the images below to enlarge

Click the images below to enlarge

Thursday, July 12, 2012
Study to Investigate the Safety and Effectiveness of TissuGlu in Abdominoplasty Surgeries Without the Use of Postoperative Drains

Cohera Medical, Inc.®, a leading innovator and developer of absorbable surgical adhesives and sealants, today announced the start of a no-drain clinical trial in 30 patients at two prominent surgery centers in Germany. The study will investigate the safety and effectiveness of TissuGlu® Surgical Adhesive in the reduction of wound drainage, post operative seroma and related complications in abdominoplasty patients without the use of drains.
Currently, most patients who undergo abdominoplasty procedures and other large flap procedures require the insertion of drains to remove fluids that accumulate under the skin at the surgical site. In some cases, drainage is inadequate and the excess fluid accumulation (seroma) requires additional procedures for removal. The drains are often painful for the patient, can lead to infection and can impact the recovery process.
TissuGlu adheres the tissue flap created during the surgical procedure to the underlying tissue, helping to reduce the space where fluids can accumulate thereby eliminating the need to use surgical drains. The ability to perform the procedure without drains would lead to a more comfortable recovery and a quicker return to normal activity for patients.
"We are excited to be part of this study utilizing TissuGlu to eliminate postoperative drains," said Dirk Richter, MD, of Wesseling, Germany. "We look forward to enrolling more patients and believe that conducting abdominoplasty procedures without the need for drains would represent a significant improvement in patient care and comfort in these procedures. The initial patients enrolled appear to be doing very well, with no seromas or complications reported in the first 48 hours after surgery."
"Initiation of this 'no drain' trial represents a major goal for TissuGlu," said Patrick Daly, President and Chief Executive Officer of Cohera Medical. "We continue to make positive patient outcomes a priority and believe the elimination of post-surgical drains is an important milestone."
Drain management and discomfort is the leading patient complaint for plastic surgery procedures according to the top plastic surgery publications. In addition, complication rates from seroma formation range from 15% to 52% following abdominoplasty procedures.
Monday, July 9, 2012
China Biologic Receives SFDA Manufacturing Approval Certificate for Human Coagulation Factor VIII
China Biologic Products, Inc., a leading fully integrated plasma-based biopharmaceutical company in China, today announced that its indirectly owned subsidiary, Shandong Taibang Biological Products Co., Ltd. ("Taibang") has received a manufacturing approval certificate from the China State Food and Drug Administration ("SFDA") for Human Coagulation Factor VIII ("FVIII"). With this certificate, the only approval remaining for Taibang's commercial production of FVIII is the SFDA's good manufacturing practice ("GMP") certification of the FVIII production line itself.
Taibang began research for FVIII in 2007 and successfully developed the technology in 2008. The Company conducted clinical trials from 2009 to 2010. In June 2010, the Company submitted required materials to Center for Drug Evaluation for approval to start manufacturing and passed on-site products verification in January 2011. The Company received official manufacturing approval certificate on June 21, 2012. FVIII will be primarily used in the treatment of hemophilia A.
Mr. David Gao, Chairman & CEO, said, "Receiving SFDA manufacturing approval represents the culmination of more than five years of work and the first technological approval of a coagulation factor product developed by Taibang. With the addition of Factor VIII, we should soon be able to offer three major categories of blood products: albumin products, immunoglobulin products and coagulation factor products. This will further strengthen our competitive position as a leading plasma-based biopharmaceutical company in China."
"We are very proud to bring FVIII to hemophilia A patients in China," continued Mr. Gao. "We believe that Human Coagulation Factor VIII addresses a critically unmet need of a large patient population and helps ease under-supply of coagulation products. Although it is not possible to specify a definitive timeframe in which we will receive GMP approval, we expect our FVIII production to be inspected by SFDA and then GMP certified by the end of 2012. Going forward, we intend to continue to develop and manufacture life-enhancing, effective, quality-assured products."
About China Biologic Products, Inc.
China Biologic Products, Inc., is a leading fully integrated plasma-based biopharmaceutical company in China. The Company's products are used as critical therapies during medical emergencies and for the prevention and treatment of life-threatening diseases and immune-deficiency related diseases. China Biologic is headquartered in Beijing and manufactures over 20 plasma-based products through its indirect majority-owned subsidiaries, Shandong Taibang Biological Products Co., Ltd. and Guiyang Dalin Biologic Technologies Co., Ltd. The Company also has an equity investment in Xi'an Huitian Blood Products Co., Ltd. The Company sells its products to hospitals and other healthcare facilities in China. For additional information, please see the Company's website www.chinabiologic.com .
Taibang began research for FVIII in 2007 and successfully developed the technology in 2008. The Company conducted clinical trials from 2009 to 2010. In June 2010, the Company submitted required materials to Center for Drug Evaluation for approval to start manufacturing and passed on-site products verification in January 2011. The Company received official manufacturing approval certificate on June 21, 2012. FVIII will be primarily used in the treatment of hemophilia A.
Mr. David Gao, Chairman & CEO, said, "Receiving SFDA manufacturing approval represents the culmination of more than five years of work and the first technological approval of a coagulation factor product developed by Taibang. With the addition of Factor VIII, we should soon be able to offer three major categories of blood products: albumin products, immunoglobulin products and coagulation factor products. This will further strengthen our competitive position as a leading plasma-based biopharmaceutical company in China."
"We are very proud to bring FVIII to hemophilia A patients in China," continued Mr. Gao. "We believe that Human Coagulation Factor VIII addresses a critically unmet need of a large patient population and helps ease under-supply of coagulation products. Although it is not possible to specify a definitive timeframe in which we will receive GMP approval, we expect our FVIII production to be inspected by SFDA and then GMP certified by the end of 2012. Going forward, we intend to continue to develop and manufacture life-enhancing, effective, quality-assured products."
About China Biologic Products, Inc.
China Biologic Products, Inc., is a leading fully integrated plasma-based biopharmaceutical company in China. The Company's products are used as critical therapies during medical emergencies and for the prevention and treatment of life-threatening diseases and immune-deficiency related diseases. China Biologic is headquartered in Beijing and manufactures over 20 plasma-based products through its indirect majority-owned subsidiaries, Shandong Taibang Biological Products Co., Ltd. and Guiyang Dalin Biologic Technologies Co., Ltd. The Company also has an equity investment in Xi'an Huitian Blood Products Co., Ltd. The Company sells its products to hospitals and other healthcare facilities in China. For additional information, please see the Company's website www.chinabiologic.com .
Friday, June 8, 2012
Study with Fibrocaps(TM) Lead product on track for market launch in 2014
LEIDEN, The Netherlands & SEATTLE, Jun 06, 2012 (BUSINESS WIRE) -- ProFibrix B.V., a leader in the development of innovative bioactive products to stop bleeding (hemostasis), today announced the start of its pivotal Phase III clinical trial with Fibrocaps (FINISH-3) in spine, liver, vascular and soft tissue surgery.
FINISH-3 is a multicenter, randomized, single-blind, controlled Phase III trial of Fibrocaps in 672 surgical patients with mild to moderate surgical bleeding. The main objectives of the study are to demonstrate superior efficacy of Fibrocaps vs. gelatin sponge within each surgical indication, and to confirm the overall safety results from the Phase II Fibrocaps trials completed in 2011.
Dr. Paul Frohna, Chief Medical Officer of ProFibrix said: "We are very pleased to announce the randomization and treatment of the first patients in the international FINISH-3 trial less than 4 months after completing our End-of-Phase II meeting with the U.S. FDA. Based on the excellent Phase II study results, the investigators are very eager to enroll their patients into our study and we remain on track for regulatory filings in the U.S. and EU in 2013. Based on the product's unique properties, and the overwhelmingly positive feedback we are receiving from the surgical community, we believe Fibrocaps should be able to capture a substantial share of the US$ 1 billion topical hemostat market."
The Fibrocaps used in the pivotal Phase III clinical trial has been manufactured by ProFibrix's commercial manufacturing partner Nova Laboratories Limited (Leicester, UK). Peter White, Managing Director of Nova Laboratories said: "It is great to be part of this exciting project and to manufacture the Phase III product and I am looking forward to our long term collaboration."
About Fibrocaps
Fibrocaps is a mixture of two essential blood clotting proteins, fibrinogen and thrombin, and is a unique dry powder topical fibrin sealant being developed to stop bleeding during or after surgery. Fibrocaps is clearly differentiated from existing liquid tissue sealants and hemostats: it is ready for immediate use, and is stable at room temperature.
About the FINISH-3 trial
FINISH-3 is a prospective, randomized (2:1), single-blind, controlled, pivotal Phase III trial of Fibrocaps vs. active control in 672 subjects undergoing spinal (n=168), liver (n=168), vascular (n=168) and soft tissue surgery (n=168). The study will be conducted at 65 sites across Europe and the U.S. Estimated completion date is May 2013.
For more details on the study, please go to http://www.clinicaltrials.gov .
About ProFibrix
ProFibrix ( www.profibrix.com ) was founded in 2004 and is headquartered in Leiden, The Netherlands, with a subsidiary in Seattle, WA, USA. The company leverages its expertise in fibrinogen technology to develop and bring to market innovative products for the hemostasis and regenerative medicine markets. Human fibrinogen plays a pivotal role in blood clotting and tissue healing. ProFibrix is led by a team with extensive commercial, clinical and scientific experience in the hemostasis field.
SOURCE: ProFibrix
Thursday, June 7, 2012
HemCon Adds Barry Starkman as CEO

HemCon Medical Technologies, Inc., an industry leader in hemorrhage control technologies and developer of lyophilized (freeze-dried) human plasma for the U.S. Army, announced today the appointment of Barry Starkman as Chief Executive Officer. Starkman joins HemCon with more than 30 years of experience in biologics and medical technology manufacturing.
"As we reorganize and restructure with the objective of successfully exiting Chapter 11, Barry is an extremely important addition to the HemCon team. His leadership and expertise will prove valuable in realizing the full potential of our lyophilized plasma program (LyP program), and in developing the optimal strategy for the Medical Device division," said Bill Wiesmann, Chairman and Co-Founder of HemCon. "Barry rounds out our executive team, working with Nick Hart who will continue his role as President and CFO."
As Chief Executive Officer, Starkman's focus will be on the execution and delivery of the LyP program through Phase 2 clinical trials and planning and executing on facility expansion to allow HemCon to enter Phase 3 clinical trials for the LyP program. He will also review the Medical Device division fully to determine the optimal means by which this business can be developed and contribute towards HemCon's overall financial performance.
Starkman's experience spans pharmaceutical products, biotech, and medical devices, matching the commercial applications for HemCon's LyP Program and Medical Devices division. His background also includes manufacturing management in the areas of facilities design, cGMP manufacturing requirements and Lean 6 Sigma applications.
Prior to joining HemCon, Starkman served as Vice President of Operations at Promega, where he was responsible for global manufacturing, planning and logistics for the $300 million organization. Starkman had previously overseen the design, construction, start-up and operation of Genentech's $450 million state-of-the-art formulation, packaging and distribution facility in Portland, serving as General Manager. Earlier in his career, Starkman worked for 24 years for Merck taking on increasing responsibility which culminated at Director of Manufacturing within Vaccine Operations.
"I am thrilled to become part of the HemCon team and bring my leadership skills to a company with breakthrough innovation in the plasma world and a proven track record of efficacious and safe medical devices which give HemCon immense potential," said Barry Starkman. "I look forward to working with the team in a timely exit from Chapter 11 and then helping HemCon realize its potential as a highly successful and viable enterprise."
HemCon Medical Technologies, Inc. ( www.hemcon.com ), founded in 2001, develops, manufactures, and markets innovative technologies for rapid delivery of plasma and hemostatic devices for the control of bleeding resulting from trauma or surgery. HemCon products are designed for use by military and civilian first responders as well as medical professionals in hospital and clinical settings where rapid supply of plasma and control of bleeding are of critical importance. HemCon is headquartered in Portland, Ore., with additional commercial operations in Ireland and the Czech Republic.
SOURCE: HemCon Medical Technologies, Inc.
Thursday, May 31, 2012
HaemoCer Instrumentation Advances Plant-sourced Hemostat Applications into Minimally Invasive Surgeries
BAYREUTH, Germany, May 31, 2012 /PRNewswire/ --BioCer Entwicklungs GmbH (BCE), a German based medical device manufacturer, announces the CE approval of its advanced HaemoCer™ Universal Applicator (UA). HaemoCer™ UA is a delivery configuration for applications of the HaemoCer™ Absorbable Polysaccharide Hemostat (APH) system in minimally invasive surgical procedures.
The HaemoCer™ UA system is a unique instrumentation for laparoscopic, ENT, spinal and other MIS procedures where precise delivery of APH particles to the bleeding site is mandatory. The delivery instrument is easily attached to the HaemoCer™ bellows dispenser and enables hemostat delivery under direct vision to the wound site for the control of capillary, venous and arteriolar bleeding. The initial HaemoCer™ 5 gram product range has also expanded to include multiple size configurations meeting the strong international market demand for HaemoCer™ in multidisciplinary procedures.
Dr. Markus Heinlein, Managing Director of BioCer, commented, "The clinical introduction of HaemoCer™ UA offers surgeons a delivery option within the expanding practice of minimally invasive surgery. Collaborating with leading laparoscopic surgeons and German engineers, our system has been designed for the controlled release of hemostatic particles. HaemoCer™ UA encompasses 3 unique advances, including a press-and-release locking system facilitating UA tip placement without uncontrolled discharge. Secondly the system features a tip design which minimizes tip blockage, both common difficulties with current particle delivery devices. Finally the new instrumentation is sold separately and the push-on coupling sheath will accommodate other devices allowing medical professionals and patients to immediately benefit from this new instrumentation while converting stocks to HaemoCer™."
BioCer Entwicklungs-GmbH, HaemoCer APH particles are manufactured utilizing a Polysaccharide Ultra-hydrophilic Resorbable Engineering (PURE) process. HaemoCer™ contains no human or animal components and is developed and manufactured in Germany. For information, distribution inquiries, and licensing options, please visit http://www.biocer-gmbh.de or email info@biocer-gmbh.de.
Sunday, May 27, 2012
Baxter Announces ADVATE Approval in China for the Treatment of Hemophilia A

Baxter International Inc. announced the approval of ADVATE[Recombinant Human Coagulation Factor VIII for injection] for the control and prophylaxis of bleeding episodes in individuals with hemophilia A (congenital factor VIII deficiency) in China by the State Food and Drug Administration (SFDA). It is estimated that more than 50,000 people in China are living with hemophilia A.
''The introduction of recombinant FVIII therapies in China offers new treatment options for hemophilia patients. The launch of ADVATE is another step in advancing hemophilia care in China,'' said Professor Yang Renchi, Institute of Hematology and Blood Diseases Hospital, Chinese Academy of Medical Sciences and Peking Union Medical College, the leading professional hematological institution providing basic medical research with clinical services in China.
''Great strides have been made in managing hemophilia, allowing people with this serious condition to live longer, more active and fulfilling lives than ever before,'' said Guan Tao, Secretary General of Hemophilia Home, the hemophilia patient organization in China. ''The availability of ADVATE will be an important milestone for people with hemophilia in China.''
ADVATE is infused directly into the bloodstream and works by temporarily raising the level of factor VIII in the bloodstream, allowing the body's blood clotting process to properly function. Extensive global use and multiple clinical trials demonstrate clinical evidence for ADVATE. With SFDA's action, ADVATE is now approved in 54 countries worldwide.
''The approval of ADVATE in China marks an important milestone for Baxter and supports our ongoing commitment to treating individuals living with hemophilia,'' said Ludwig Hantson, Ph.D., president of Baxter's BioScience business.
Baxter continues to work closely with the Chinese hemophilia community, including both patients and treaters, to provide access to care for this life-saving, life-sustaining therapy. In 2010, Baxter cooperated with the Ministry of Health to set up a ''Hemophilia Disease Management System,'' China's first nationwide hemophilia patient registration and management system integrating diagnosis and treatment information. In recent years, Baxter has donated more than five million IUs of hemophilia products to Chinese patients and has provided a number of resources to raise awareness of the disease.
About ADVATE
ADVATE [Antihemophilic Factor (Recombinant) Plasma/Albumin-Free Method] was initially approved by the FDA in July 2003 for control and prevention of bleeding episodes in adults and children (0-16 years) with hemophilia A. ADVATE is a full-length (derived from the complete FVIII gene) recombinant FVIII product that is processed without any blood-based additives. Because no blood derived components are added at any stage of the manufacturing process, the potential risk of transmitting pathogens that may be carried in blood-based additives is eliminated. There have been no confirmed reports of transmission of HIV, HBV or HCV with rFVIII therapies.
ADVATE is approved in the United States, Canada, 27 countries in the European Union, Argentina, Australia, Brazil, Chile, China, Colombia, Croatia, Hong Kong, Iceland, Iraq, Japan, Macau, Malaysia, New Zealand, Norway, Panama, Puerto Rico, Serbia, Singapore, South Korea, Suriname, Switzerland, Taiwan, Uruguay and Venezuela.
In the United States, ADVATE [Antihemophilic Factor (Recombinant) Plasma/AlbuminFree Method] is also indicated for routine prophylaxis to prevent or reduce the frequency of bleeding episodes in adults and children (0-16 years) with hemophilia A. ADVATE is the only antihemophilic factor approved in the United States for prophylactic use in both adults and children. ADVATE is not indicated for the treatment of von Willebrand disease.
FastSeal(R) Bioabsorbable Vascular Access Closure System
Vascular Closure Systems, Inc, First in Human (FIH) Use of the FastSeal(R) Bioabsorbable Vascular Access Closure System -- 100% Success Rate, Including Patients with Challenging Anatomy and Vessel Condition.
The company is pleased to announce that after receiving Ethical Committee (EC) and Ministry of Health (MOH) approval, for the first human use of the company's FastSeal(R) Bioabsorbable Vascular Access Closure System, Phase I of the First in Human (FIH) clinical testing has been completed, with 100% successful results, and no adverse effects, including post procedural discomfort. The average Time to Hemostasis (TTH) was less than one minute. The clinical cases included vessels that were normal, diseased, severely fibrosed and with calcified plaque.
The initial series of human clinical cases were performed on Percutaneous Coronary Intervention (PCI) patients, by Prof. Alessandro Bortone of the Policlinico di Bari, University of Bari School of Medicine (IT). Phase II testing will begin within the next two weeks.
The detailed FIH test results will be presented at multiple upcoming medical conferences.
The company is planning to begin International commercialization (pending regulatory approval) during the fourth quarter of this year.
About FastSeal(R)
Our Bioabsorbable Vascular Access Closure System is packaged and used as a single piece unit, with no assembly required prior to use, and no separate deployment device is needed to be inserted into the puncture site. Simply insert the FastSeal(R) system into the hub of the procedural introducer sheath, and advance the attached plunger. The system design enables hemostasis within less than a minute after the non-collagen sealing element has been deployed. Our system doesn't require the use of a specific type or brand of vascular introducer sheath, and is compatible with any commercialized vascular introducer sheath with a useable length of between 10 to 12 cm. Once the sealing element has been deployed, no external compression is required. The inner vessel section of the sealing element is absorbed within 10 to 14 days. The remainder of the sealing element is completely absorbed within 21 days. The FastSeal(R) system has the ability to be removed after deployment (if desired), without causing trauma to the vessel or requiring a surgical intervention.
Subscribe to:
Posts (Atom)